Indonesia Medical Device Registration & Regulatory Services
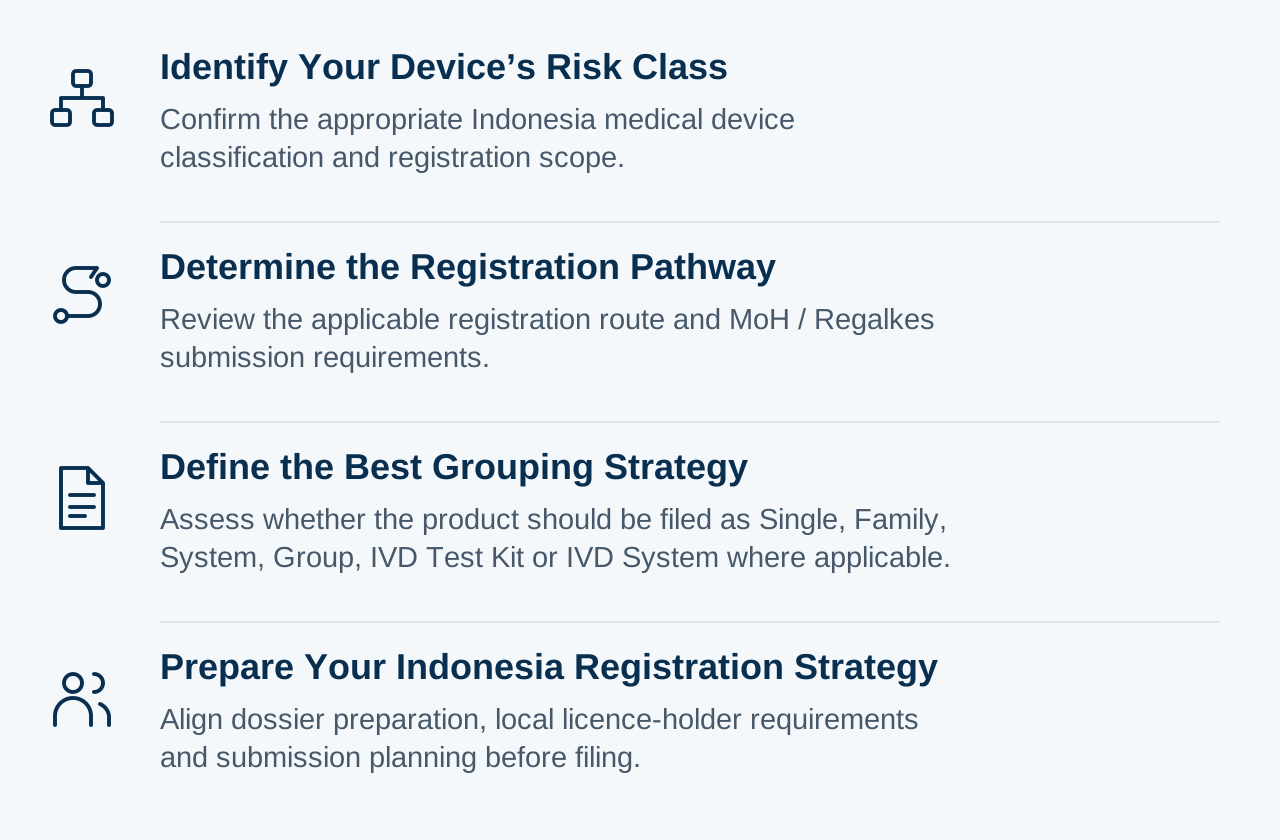
MediPass helps you navigate Indonesian Ministry of Health requirements with clarity — from product classification, grouping and local licence-holder readiness to dossier preparation, Regalkes coordination and post-market compliance.
Classification & Filing
Risk Class A-D, with grouping strategies planning
Registration Pathways
Submission strategy and Regalkes coordination.
Local Registrant
Import coordination
Post-Market Obligations
Complaint Handling, Adverse Event Reporting, FSCA & Recall
ASEAN Expansion
Indonesia & Southeast Asia
Strategy
ASEAN EXPANSION STRATEGY
One Regional Partner
for Foreign Manufacturers
The ASEAN Medical Device Directive (AMDD) supports regulatory harmonisation across Southeast Asia, but each market maintains its own approval process and local requirements. In Indonesia, medical devices are regulated by the Indonesia Ministry of Health (MoH).
For foreign manufacturers, MediPass supports Indonesia market entry through classification, registration planning, local importer coordination and post-market compliance. We also serve as one regional partner for broader ASEAN expansion across Singapore, Malaysia, Thailand, and Vietnam markets.
Indonesia Market Snapshot
Key demographic, economic, and medical device registration indicators relevant to Indonesia market entry.
Country Overview
Selected macro indicators based on 2025 data
Population
Median Age
GDP
GDP per Capita
Indonesia MoH
Medical Device Product Applications Processed by Thai FDA (FY2022–FY2025)
Domestic Medical Device
Imported Medical Devices
Domestic PKRT Products
Imported PKRT Products
Regulatory Overview
Medical Devices in Indonesia
Regulated by the Indonesian Ministry of Health under Ministry of Health Regulation No. 62 of 2017 and Regulation No. 11 of 2025. Medical devices, including IVDs, are classified into Classes A–D based on risk. Foreign manufacturers must appoint a licensed Indonesian distributor/local licence holder, who submits the product application through the Regalkes system.
Registration Process
Determine
Device Status
Classify
Class A/B/C/D
Filing Plan
Grouping
Prepare
CSDT Dossier
Submit
Regalkes System
Registration Pathways
In Indonesia, all imported devices require an appointed Indonesian licence holder, a legalised or apostilled LOA, a valid CFS or marketing authorisation, QMS evidence and a product dossier. Classes B–D generally follow the same document-review route; the risk class mainly affects the review depth.
Class 1A
Low-risk devices. Standard registration takes up to 21 working days. Products included in the MoH list may use the simplified Notification Route, taking up to 12 working days.
Class 2B
Low–moderate-risk devices. Applications undergo standard dossier review, with a maximum service period of 36 working days.
Class 2C
Moderate–high-risk devices. The same core dossier and 36-working-day service period generally apply, specified products may require testing by an Indonesian accredited or MoH-designated laboratory.
Class 3D
Highest level of review, with a maximum service period of 55 working days. A complete technical, clinical and risk-management dossier is required; Specified products may require testing by an Indonesian accredited or MoH-designated laboratory.
Grouping Strategy for Efficient Registration Planning
In Indonesia, medial devices may be classified as a Single Device, Family, System, Group, IVD Test Kit or IVD Cluster. Early grouping helps align the risk class, model scope, labels and technical dossier while avoiding duplicate / split applications.
· Permitted variants · One Product Owner · One common intended purpose
Single
A medical device from one product owner, identified by a proprietary or brand name, with a specific intended purpose, with permitted variations such as package size or quantity where the criteria allow.
Family
A collection of medical devices from the same product owner and risk classification, sharing a common intended purpose, common design and manufacturing process, and only permissible variants (size, length, diameter, colour, flexibility, memory capacity, sterilization method and etc.)
System
Compatible medical devices and accessories intended to work together for one common medical purpose. Their labels, IFUs or catalogues must establish their use as a system. Third-party components may be included where the system owner assumes responsibility and provides supporting compatibility evidence.
Group
Related systems with the same brand, common intended purpose and shared design and manufacturing characteristics, with permitted component variants.
IVD Test Kit
Compatible reagents or articles used together for one specific diagnostic purpose under a common test-kit name. An analyser is not included and should normally be handled as part of an IVD system. Replacement reagents may be supplied separately within the approved scope.
IVD System
Eligible IVD reagents or articles from the same product owner. Products must share the same risk class, test methodology and recognized cluster category. Class C or D IVDs cannot use this grouping route.
Get Your Free Indonesia Market Assessment
Submit your product details and let MediPass recommend the most efficient registration strategy — helping you clarify classification, grouping, local regulatory requirements and market-entry planning in Indonesia.
Need help with classification, grouping strategy, or registration pathway?
*Accepted file types include IFUs, Brochures, or existing Approval documents. Please attach to help us assess your product.
Initial assessment within 1 business day
| Indonesia MoH Medical Device Fee Overview | ||
|---|---|---|
| Device Classification | MoH Fee | Indicative Review Time |
| Class 1A | IDR 1,500,000 | 1 month |
| Class 2B / 2C | IDR 3,000,000 | 2 months |
| Class 3D | IDR 5,000,000 | 2–3 months |
| Indonesia MoH PKRT Fee Overview | ||
|---|---|---|
| PKRT Classification | MoH Fee | Indicative Review Time |
| Class I | IDR 1,000,000 | 45 days |
| Class II | IDR 2,000,000 | 2 months |
| Class III | IDR 3,000,000 | 3 months |